Cadrenal Therapeutics (CVKD) – 2Q26 Reported With Review Of New “Three Pillars” Strategy


Monday, August 17, 2026

Robert LeBoyer, Senior Vice President, Equity Research Analyst, Biotechnology, Noble Capital Markets, Inc.

Refer to the full report for the price target, fundamental analysis, and rating.

2Q26 Reported With Review Of New Strategy and Product Data. Cadrenal reported a 2Q26 loss of $3.3 million, or $(1.14) per share. The company modified its strategy and plans to develop its products through collaborations, out-licensing agreements, and non-dilutive grants to conserve capital resources. On June 30, 2026, cash and cash equivalents were $4.2 million, excluding proceeds from the private placement completed July 1. The private placement raised about $3.0 million, with warrants that could raise another $5.8 million upon exercise.

The Pipeline Has Been Reorganized Into “Three Pillars”.  The company has divided the pipeline into products for Cardiac Acute Critical Care, Orphan Diseases, and Post-Operative Care. These divisions emphasize how the products can address important needs before and after cardiac surgery, as well as for patient populations with few options.


Get the Full Report

Equity Research is available at no cost to Registered users of Channelchek. Not a Member? Click ‘Join’ to join the Channelchek Community. There is no cost to register, and we never collect credit card information.

This Company Sponsored Research is provided by Noble Capital Markets, Inc., a FINRA and S.E.C. registered broker-dealer (B/D).

*Analyst certification and important disclosures included in the full report. NOTE: investment decisions should not be based upon the content of this research summary. Proper due diligence is required before making any investment decision. 

Eledon Pharmaceuticals (ELDN) – 2Q26 Reported With Several Tegoprubart Trial Updates


Friday, August 14, 2026

Robert LeBoyer, Senior Vice President, Equity Research Analyst, Biotechnology, Noble Capital Markets, Inc.

Refer to the full report for the price target, fundamental analysis, and rating.

2Q Financial Results Were Within Expectations. Eledon reported a 2Q26 loss of $31.6 million or $(0.27) per share. The Operating Loss of $22.9 million was close to our estimate of $22.4 million, before a charge of $9.6 million for Changes In The Fair Value of Warrant Liabilities. The Net Loss excluding the non-cash charge would have been $22 million. Cash balance on June 30, 2026 was $ $88.8 million.

Preparations For A Global Phase 3 Trial In Kidney Transplantation Continue. During 2Q26, an End-Of-Phase 2 meeting was held with the FDA to discuss the Phase 3 trial design and requirements for a BLA submission. The Phase 3 trial has been designed to test tegoprubart against tacrolimus to prevent kidney transplant rejection. It is scheduled to begin in late 2026 with a target enrollment of about 600 patients. The Primary Endpoint will be non-inferiority based on a composite of BPAR (biopsy-proven acute rejection), graft loss, and death.


Get the Full Report

Equity Research is available at no cost to Registered users of Channelchek. Not a Member? Click ‘Join’ to join the Channelchek Community. There is no cost to register, and we never collect credit card information.

This Company Sponsored Research is provided by Noble Capital Markets, Inc., a FINRA and S.E.C. registered broker-dealer (B/D).

*Analyst certification and important disclosures included in the full report. NOTE: investment decisions should not be based upon the content of this research summary. Proper due diligence is required before making any investment decision. 

Release – Cadrenal Therapeutics Reports Second Quarter 2026 Financial Results; Provides a Corporate Update on the Cardiac Acute Critical Care Franchise and the Strategic Partnering Process

Primary Logo

Research News and Market Data on CVKD

Late-breaking Phase 2 CAD-1005 data presented at ISTH support a late-stage portfolio organized around preoperative safety, orphan regulatory acceleration, and postoperative shielding

PONTE VEDRA, Fla., Aug. 13, 2026 (GLOBE NEWSWIRE) — Cadrenal Therapeutics, Inc. (Nasdaq: CVKD), a late-stage biopharmaceutical company advancing specialized therapies for critical care cardiology and orphan cardiovascular conditions, today reported financial results for the second quarter ended June 30, 2026, and provided a corporate update on the launch of a structured strategic partnering process following significant clinical, regulatory, and portfolio progress across its Cardiac Acute Critical Care Franchise.

“The second quarter and subsequent weeks marked an important milestone for Cadrenal,” said Quang X. Pham, Chairman and Chief Executive Officer of Cadrenal Therapeutics. “Our objective is to convert clinical and regulatory progress into strategic value while remaining capital disciplined. We are also pursuing multiple nondilutive grants to advance our programs. With CAD-1005, frunexian, and tecarfarin addressing complementary acute, critical care, and orphan cardiovascular opportunities, and CAD-2000 extending the 12-LOX platform to treat chronic conditions, we believe Cadrenal offers a differentiated pipeline that addresses critical unmet needs for high-risk patients.”

“Having achieved regulatory guidance on the Phase 3 path for CAD-1005, we delivered a late-breaking presentation to the global thrombosis community on Phase 2 data for CAD-1005, which demonstrated an absolute reduction of more than 25% in thrombotic events. Our portfolio, now organized into a Cardiac Acute Critical Care Franchise spanning pre-operative safety, orphan regulatory acceleration, and post-operative shielding, is structured to demonstrate to prospective partners how our assets can address multiple high-value needs across the cardiac surgery continuum.”

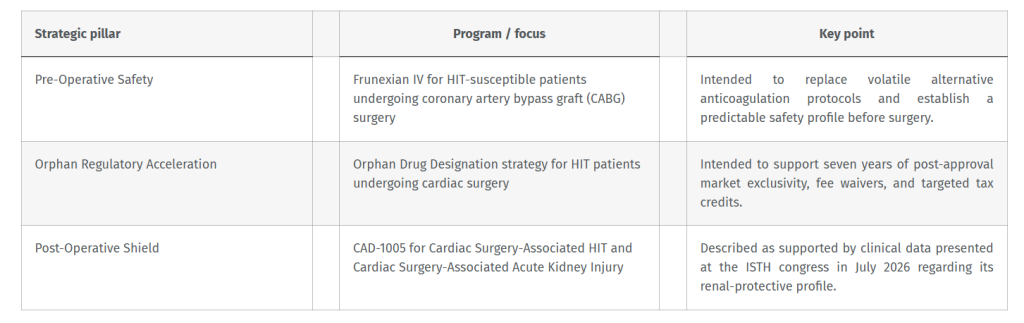
The strategic alignment of the Cardiac Acute Critical Care (CACC) Franchise organizes Cadrenal’s portfolio into three commercial pillars.

Recent Highlights

  • Presented late-breaking Phase 2 data on CAD-1005 at the International Society of Thrombosis and Hemostasis (ISTH) 2026 Congress in Paris. The randomized, blinded, placebo-controlled study demonstrated an absolute reduction of more than 25% in thrombotic events when CAD-1005 was added to standard anticoagulant therapy, with a favorable safety profile and renal-protective baseline.
  • Launched a structured partnering process to explore development, licensing, and commercialization transactions for CAD-1005, frunexian, and tecarfarin. Cadrenal is pursuing a capital-efficient model focused on strategic out-licensing, portfolio monetization, and commercial co-development rather than independently funding large late-stage clinical trials.
  • Solidified a multi-indication strategy for CAD-1005 in heparin-induced thrombocytopenia (HIT) and Cardiac Surgery-Associated Acute Kidney Injury (CSA-AKI). The Company believes that the shared cardiac intensive care infrastructure, intravenous administration, and complementary thrombotic and inflammatory mechanisms may create a differentiated critical care asset package for prospective partners.
  • Submitted a request to the U.S. Food and Drug Administration on July 8, 2026, for Rare Pediatric Disease Designation for tecarfarin for the prevention of life-threatening blood clots inside coronary artery aneurysms in children with Kawasaki Disease.
  • Signed an agreement with a global firm that has collectively secured over $500 million in non-dilutive funding for life science companies.
  • Continued advancing CAD-2000, a highly selective, orally bioavailable preclinical 12-lipoxygenase (12-LOX) inhibitor designed for chronic cardiorenal inflammatory and thrombotic indications, as a potential follow-on companion to the Company’s intravenous acute care platform.
  • Signed a private placement that generated approximately $3.0 million in gross proceeds, with up to approximately $5.8 million in additional gross proceeds if the associated warrants are exercised in full for cash. There can be no assurance that any warrants will be exercised.

Second Quarter 2026 Financial Highlights

Research and development expenses for the quarter ended June 30, 2026, were $0.7 million compared to $1.1 million for the same period in 2025. General and administrative expenses were $2.6 million compared to $2.7 million for the same period in 2025. Total operating expenses were $3.3 million compared to $3.7 million for the same period in 2025. Cadrenal reported a net loss of $3.3 million for the quarter ended June 30, 2026, compared to $3.7 million for the same period in 2025.

As of early August 2026, Cadrenal had approximately $4.2 million in cash and cash equivalents. Based on its current operating plan, the Company believes these resources are expected to fund operations through the first quarter of 2027. The Company’s existing cash resources are not sufficient to advance its product candidates to clinical trial readiness or to commence and complete any clinical trials. Cadrenal does not plan to commence a clinical trial unless funding sufficient to complete that trial is in place, which may include capital raised through strategic partnerships, out-licensing agreements, non-dilutive grants, equity or debt financing, or a combination of these sources.

About Cadrenal Therapeutics, Inc.

Cadrenal Therapeutics, Inc. is a late-stage biopharmaceutical company advancing specialized therapies for critical care cardiology and orphan cardiovascular conditions. The Company’s pipeline includes CAD-1005, tecarfarin, and frunexian. CAD-1005 is a novel investigational therapeutic in development for the treatment of heparin-induced thrombocytopenia (HIT) and Cardiac Surgery-Associated Acute Kidney Injury (CSA-AKI). CAD-1005 is designed to selectively inhibit 12-lipoxygenase (12-LOX), an enzyme central to platelet immune activation and thrombo-inflammatory signaling in HIT. CAD-1005 is intended to be used alongside existing standards of care and is being developed to address the underlying biological mechanisms that drive disease progression. CAD-1005 has an Orphan Drug Designation (“ODD”) from the U.S. Food and Drug Administration (“FDA”) for prophylaxis of thrombosis in patients with HIT, FDA Fast Track designation for the treatment and prevention of HIT, and an orphan designation from the European Medicines Agency for the treatment of platelet-activating factor 4 disorders. Second-generation 12-LOX oral therapeutics (CAD-2000) are also in development for chronic indications.

The Company’s broader pipeline includes tecarfarin, a late-stage oral vitamin K antagonist designed to prevent heart attacks, strokes, and deaths from blood clots in patients requiring chronic anticoagulation, including those with end-stage kidney disease and atrial fibrillation, those with left ventricular assist devices, and potentially those with Kawasaki disease (KD), an acute, self-limited, febrile illness that primarily affects children under 5 years old and is the leading cause of acquired heart disease in developed countries. The Company recently submitted a request for Rare Pediatric Disease Designation (RPDD) to the FDA for tecarfarin for “Prevention of the Formation of Life-Threatening Blood Clots Inside Coronary Artery Aneurysms in Children with Kawasaki Disease”. Tecarfarin has also received Orphan Drug and Fast Track designations from the FDA.

For more information, visit https://www.cadrenal.com/ and connect with the Company on LinkedIn.

Safe Harbor

Any statements in this press release about future expectations, plans, and prospects, as well as any other statements regarding matters that are not historical facts, may constitute “forward-looking statements.” The words “anticipate,” “believe,” “continue,” “could,” “estimate,” “expect,” “intend,” “may,” “plan,” “potentially,” “predict,” “project,” “should,” “target,” “will,” “would” and similar expressions are intended to identify forward-looking statements, although not all forward-looking statements contain these identifying words. These statements include, without limitation, statements regarding the Company converting clinical and regulatory progress into strategic value while remaining capital disciplined; pursuing multiple nondilutive grants to advance the Company’s programs; Cadrenal offering a differentiated pipeline that addresses critical unmet needs for high-risk patients; the CACC Franchise demonstrating to prospective partners how the Company’s assets can address multiple high-value needs across the cardiac surgery continuum; frunexian IV replacing volatile alternative anticoagulation protocols for HIT-susceptible patients undergoing CABG surgery and establishing a predictable safety profile; ODD for HIT supporting seven years of post-approval market exclusivity, fee waivers and targeted tax credits; CAD-1005 serving as a post-operative shield for Cardiac Surgery-Associated HIT and CSA-AKI; the Company’s ability to enter into development, licensing, and commercialization transactions for CAD-1005, frunexian, and tecarfarin; the shared cardiac intensive care infrastructure, intravenous administration, and complementary thrombotic and inflammatory mechanisms creating a differentiated critical care asset package for prospective partners; the FDA’s ultimate decision regarding the Company’s request for RPDD for tecarfarin for the prevention of life-threatening blood clots inside coronary artery aneurysms in children with Kawasaki Disease; CAD-2000 potentially being a follow-on companion to the Company’s intravenous acute care platform; the potential exercise of the warrants issued in the Company’s private placement resulting in gross proceeds of up to $5.8 million; the Company’s belief that its resources will fund operations through the first quarter of 2027; the Company securing funding sufficient to complete a trial, which may include capital raised through strategic partnerships, out-licensing agreements, non-dilutive grants, equity or debt financing, or a combination of these sources; and tecarfarin, a late-stage oral vitamin K antagonist designed to prevent heart attacks, strokes, and deaths from blood clots in patients requiring chronic anticoagulation, potentially treating patients with Kawasaki disease. Actual results may differ materially from those indicated by such forward-looking statements as a result of various important factors, including the Company’s ability to advance its programs to clinical trial readiness; the Company’s ability to enter into development, licensing, and commercialization transactions for CAD-1005, frunexian, and tecarfarin; the Company’s ability to secure nondilutive grants to advance its programs; and the other risk factors described in the Company’s Annual Report on Form 10-K for the year ended December 31, 2025, and the Company’s subsequent filings with the Securities and Exchange Commission, including subsequent periodic reports on Quarterly Reports on Form 10-Q and Current Reports on Form 8-K. Any forward-looking statements contained in this press release speak only as of the date hereof and, except as required by federal securities laws, the Company specifically disclaims any obligation to update any forward-looking statement, whether as a result of new information, future events, or otherwise.

For more information, please contact:

Lytham Partners, LLC
Robert Blum, Managing Partner
602-889-9700
[email protected]

View full release here.

Release – Eledon Pharmaceuticals Reports Second Quarter 2026 Financial Results and Recent Business Highlights

eledon logo

Research News and Market Data on ELDN

August 13, 2026

PDF Version

Successful End-of-Phase 2 meeting with FDA supports advancement of tegoprubart into a global Phase 3 kidney transplantation trial, on track to initiate in late 2026

Long-term Phase 2 BESTOW data presented at ATC 2026 demonstrated sustained higher kidney function and improved patient-reported outcomes with tegoprubart compared with tacrolimus

Updated islet cell transplantation data presented at ADA 2026 showed 100% insulin independence in all 12 patients with type 1 diabetes

Cash, cash equivalents and short-term investments of $88.8 million as of June 30, 2026

IRVINE, Calif., Aug. 13, 2026 (GLOBE NEWSWIRE) — Eledon Pharmaceuticals, Inc. (“Eledon”) (Nasdaq: ELDN) today reported its second quarter 2026 operating and financial results and provided recent business highlights.

“During the first half of the year, we made meaningful progress establishing the regulatory framework for our planned Phase 3 kidney transplantation program, which we expect to initiate later this year,” said David-Alexandre C. Gros, M.D., Chief Executive Officer of Eledon. “The continued strength of our clinical data, including sustained long-term kidney function in BESTOW and the compelling results from the UChicago Medicine islet cell transplantation study, reinforces the potential of tegoprubart to improve outcomes across multiple transplant settings. We look forward to advancing our Phase 3 program and expanding the clinical evidence for tegoprubart across additional transplant indications.”

Second Quarter 2026 Business Highlights

Kidney Transplantation

  • Completed a successful End-of-Phase 2 meeting with the U.S. Food and Drug Administration, establishing the regulatory framework for the planned Phase 3 trial of tegoprubart in kidney transplantation. The global trial is on track to initiate in late 2026 and enroll approximately 600 patients, with a primary endpoint of non-inferiority versus tacrolimus at 52 weeks based on a composite of biopsy-proven acute rejection (BPAR), graft loss and death.
  • Presented new long-term data from the Phase 2 BESTOW clinical program at the American Transplant Congress (ATC) in June 2026, demonstrating sustained higher kidney function in kidney transplant patients treated with tegoprubart compared with tacrolimus, the current standard-of-care immunosuppression therapy. At 18 months, the eGFR curves showed a statistically significant separation (p<0.05), with mean eGFR approximately 12 mL/min/1.73 m² higher for tegoprubart compared with tacrolimus (approximately 74 vs. 61 mL/min/1.73 m²). No BPAR events were observed in tegoprubart-treated patients after the first six months post-transplant, compared with seven BPAR events (9.4% of tacrolimus-treated patients) reported in the tacrolimus arm. Patient-reported outcomes at 52 weeks favored tegoprubart, with statistically significant improvements versus tacrolimus on two validated measures of symptom burden. Long-term data from the BESTOW extension trial also demonstrated favorable long-term safety and tolerability.
  • Treated the first two patients in an investigator-initiated study of kidney transplant tolerance induction at Massachusetts General Hospital.
  • Entered into a strategic partnership with Natera, Inc., a global leader in cell-free DNA testing and precision medicine, to incorporate Natera’s Prospera kidney transplant assessment test as the exclusive donor-derived cell-free DNA (dd-cfDNA) monitoring assay in Eledon’s planned Phase 3 kidney transplantation trial.

Islet Cell Transplantation

  • Presented updated data from the University of Chicago Medicine investigator-initiated islet cell transplantation study at ADA 2026. All 12 patients with T1D achieved insulin independence and HbA1c below 6.5%, with a mean most recent HbA1c of approximately 5.4% and no severe hypoglycemic episodes post-transplant. Tegoprubart demonstrated stable islet graft function through a maximum follow-up of 22 months and was generally well tolerated, with no evidence of nephrotoxicity, hypertension or neurotoxicity.

Anticipated Upcoming Milestones

The Company anticipates the following milestones in 2026 and over the next 12 months:

  • Initiate Phase 3 clinical trial evaluating tegoprubart in kidney transplantation in late 2026.
  • Support the initiation of an investigator-led study evaluating tegoprubart for the prevention of organ rejection in patients with renal dysfunction receiving an islet cell transplant in 2026.
  • Initiate company-sponsored, registration path study evaluating tegoprubart in islet cell transplantation.
  • Support the initiation of an investigator-led study evaluating tegoprubart for the prevention of organ rejection in patients receiving a de novo liver transplant.
  • Receive FDA regulatory guidance on the path to market for tegoprubart in xenotransplantation.

Second Quarter 2026 Financial Results

Cash, cash equivalents and short-term investments totaled $88.8 million as of June 30, 2026, compared to $133.3 million as of December 31, 2025. The Company expects current cash, cash equivalents and short-term investments to fund operations into the second quarter of 2027.

Research and development (R&D) expenses for the second quarter of 2026 were $18.2 million, including $2.1 million of non-cash stock-based compensation expense, compared to $20.3 million for the comparable period in 2025, including $1.1 million of non-cash stock-based compensation expense.

General and administrative (G&A) expenses for the second quarter of 2026 were $4.6 million, including $1.1 million of non-cash stock-based compensation expense, compared to $4.5 million for the comparable period in 2025, including $1.6 million of non-cash stock-based compensation expense.

Net loss for the second quarter of 2026 was $31.6 million, or $0.27 per basic common share, compared to a net loss of $11.2 million, or $0.13 per basic common share, for the comparable period in 2025. Net loss in the second quarter of 2026 included a non-cash loss of $9.6 million from changes in the fair value of warrant liabilities, while the 2025 net loss included a non-cash gain of $12.3 million from such changes. Excluding the non-cash items related to changes in the fair value of warrant liabilities, Eledon would have recorded a net loss of $22.0 million for the three months ended June 30, 2026, and $23.5 million for the three months ended June 30, 2025.

About Eledon Pharmaceuticals and tegoprubart

Eledon Pharmaceuticals, Inc. is a clinical stage biotechnology company that is developing immune-modulating therapies for the management and treatment of life-threatening conditions. The Company’s lead investigational product is tegoprubart, an anti-CD40L antibody with high affinity for the CD40 Ligand, a well-validated biological target that has broad therapeutic potential. The central role of CD40L signaling in both adaptive and innate immune cell activation and function positions it as an attractive target for non-lymphocyte depleting, immunomodulatory therapeutic intervention. The Company is building upon a deep historical knowledge of anti-CD40L biology to conduct preclinical and clinical studies in kidney allograft transplantation, xenotransplantation, islet cell transplantation, liver transplantation and amyotrophic lateral sclerosis (ALS). Eledon is headquartered in Irvine, California. For more information, please visit the Company’s website at www.eledon.com.

Follow Eledon Pharmaceuticals on social media: LinkedIn; X

Forward-Looking Statements

This press release contains forward-looking statements that involve substantial risks and uncertainties. Any statements about the company’s future expectations, plans and prospects, including statements about planned clinical trials, the development of product candidates, expected timing for initiation of future clinical trials, expected timing for receipt of data from clinical trials, the company’s capital resources and ability to finance planned clinical trials, as well as other statements containing the words “believes,” “anticipates,” “plans,” “expects,” “estimates,” “intends,” “predicts,” “projects,” “targets,” “looks forward,” “could,” “may,” and similar expressions, constitute forward-looking statements within the meaning of the Private Securities Litigation Reform Act of 1995. Forward-looking statements are inherently uncertain and are subject to numerous risks and uncertainties, including: our short operating history and shifts in our business strategy; our operating losses since inception; our need for additional funding to develop our lead drug candidate and our ability to secure additional funding on acceptable terms or at all; the impact of issuances of our common stock, including the possibility of dilution or a decline in our stock price; our ability to successfully develop our product candidates; unfavorable global economic and financial market conditions; the regulatory environment of our business and our ability to obtain required regulatory approvals; results of non-clinical studies and clinical trials, and risks that non-clinical studies or early clinical trials may not be predictive of results of later-stage clinical trials; delays or difficulties in enrollment of patients in clinical trials; our ability to attract and retain our executives and key employees; legislation of the pharmaceutical and healthcare industries; cybersecurity and data privacy risks; the ability of our products to achieve marketing approval; competition in our industry; our ability to obtain insurance coverage; our dependence on contract research organizations; our ability to protect our intellectual property; public health crises; our ability to maintain proper and effective internal control over financial reporting and other risks disclosed in our Annual Report on Form 10-K for the year ended December 31, 2025, filed with the Securities and Exchange Commission on March 19, 2026. Actual results may differ materially from those indicated by such forward-looking statements as a result of various factors. These risks and uncertainties, as well as other risks and uncertainties that could cause the company’s actual results to differ materially from the forward-looking statements contained herein, are discussed in our Annual Report on Form 10-K, and other filings with the U.S. Securities and Exchange Commission, which can be found at www.sec.gov. Any forward-looking statements contained in this press release speak only as of the date hereof and not as of any future date, and the company expressly disclaims any intent to update any forward-looking statements, whether as a result of new information, future events or otherwise.

Investor Contact:

Stephen Jasper
Gilmartin Group
(858) 525 2047
[email protected]

Media Contact:

Jenna Urban
CG Life
(212) 253 8881
[email protected]

ELDN

Unicycive Therapeutics (UNCY) – 2Q26 Reported As OLC Moving Forward With FDA Manufacturing Inspection


Thursday, August 13, 2026

Robert LeBoyer, Senior Vice President, Equity Research Analyst, Biotechnology, Noble Capital Markets, Inc.

Refer to the full report for the price target, fundamental analysis, and rating.

OLC Is Moving Forward. Unicycive reported a 2Q26 loss of $1.7 million, or $(0.06) per share. The Operating Loss of $10.1 million was offset by $8.0 million in Change In Fair Value Of Warrant Liabilities, leading to a Net Loss To Common Shareholders of $1.7 million. Importantly, the FDA has given written notice of facility inspection to one of the OLC third-party manufacturers. Assuming the inspection results are positive, Unicycive will be able to resubmit its NDA for OLC. Cash and equivalents on June 30, 2026, were $61.4 million.

The Third-Party Inspection Could Complete The Missing Part Of The NDA. In June 2026, Unicycive received a CRL (Complete Response Letter) to its NDA for OLC. The stated reason was that the required FDA inspection of one of its third-party manufacturing vendors had not been performed. The notification of an inspection is good news that could allow the NDA to be resubmitted.


Get the Full Report

Equity Research is available at no cost to Registered users of Channelchek. Not a Member? Click ‘Join’ to join the Channelchek Community. There is no cost to register, and we never collect credit card information.

This Company Sponsored Research is provided by Noble Capital Markets, Inc., a FINRA and S.E.C. registered broker-dealer (B/D).

*Analyst certification and important disclosures included in the full report. NOTE: investment decisions should not be based upon the content of this research summary. Proper due diligence is required before making any investment decision. 

Release – Unicycive Therapeutics Announces Second Quarter 2026 Financial Results and Provides Business Update

Research News and Market Data on UNCY

August 12, 2026 7:19am EDT 

Download as PDF

– Company expects to resubmit New Drug Application (NDA) for oxylanthanum carbonate (OLC) assuming completion of successful inspection of third-party manufacturing vendor

– FDA has assigned a facility inspection to third-party manufacturing vendor of OLC

– As of June 30, 2026, unaudited cash, cash equivalents, and marketable securities totaled $61.4 million, with expected runway into 2027

MOUNTAIN VIEW, Calif., Aug. 12, 2026 (GLOBE NEWSWIRE) — Unicycive Therapeutics, Inc. (Nasdaq: UNCY), a clinical-stage biotechnology company developing therapies for patients with kidney disease, today announced its financial results for the second quarter ended June 30, 2026, and provided a business update.

“We are focused on securing approval of oxylanthanum carbonate (OLC) and remain confident in the efficacy and safety of OLC and in its potential to improve care for patients with hyperphosphatemia on dialysis,” said Shalabh Gupta, M.D., Chief Executive Officer of Unicycive. “The latest update from our third-party manufacturing vendor is that the U.S. Food and Drug Administration (FDA) has assigned a facility inspection. This marks a positive step forward, and our dialogue with the FDA on OLC labeling and packaging has been productive and continuous throughout this process. We are optimistic about a successful inspection of the third-party manufacturing facility, which would enable us to promptly resubmit the NDA. In the meantime, we are well positioned to launch OLC as quickly as possible following potential approval, and we are utilizing this time to continue to expand market awareness in preparation for the commercial success of OLC.”

Key Highlights & Upcoming Milestones

  • In June, the Company received a Complete Response Letter (CRL) from the FDA regarding the resubmitted NDA for OLC for the treatment of hyperphosphatemia in patients with chronic kidney disease on dialysis. The CRL cites the same third-party manufacturing deficiencies identified in a previous CRL issued in June 2025. The FDA has not raised any concerns regarding clinical efficacy or safety data, and no additional data was requested from the Company.
  • The Company’s third-party vendor has received written notification from the FDA that the facility inspection has been assigned, and the Company plans to provide an update following completion of the FDA inspection.
  • In preparation for the potential launch of OLC, the Company continues to advance its commercial readiness initiatives. Unicycive is focused on optimizing patient access across all reimbursement settings and plans to support patients with dedicated access and reimbursement services through its UniSource™ reimbursement hub.
  • The Company will also engage with the patient and clinical community at several medical meetings during the third quarter, including the 51st Annual American Association of Kidney Patients National Patient Meeting (September 11–13, Little Rock, Arkansas) and the 2026 Renal Healthcare Association Annual Conference (September 23–26, Savannah, Georgia).


Financial Results for the Quarter Ended June 30, 2026

Research and Development (R&D) expense was $2.8 million for the quarter ended June 30, 2026, compared to $1.8 million for the three months ended June 30, 2025. The increase was primarily driven by a $0.9 million increase in non-cash stock-based compensation, and an increase in consulting and professional fees of $0.1 million.

General and Administrative (G&A) expense was $7.4 million for the quarter ended June 30, 2026, compared to $5.2 million for the three months ended June 30, 2025. The increase was primarily driven by a $1.4 million increase in non-cash stock-based compensation as well as an increase of $0.3 million in other labor costs. There was also an increase of $0.4 million related to commercial launch preparation.

Other income (expense) was $8.4 million for the quarter ended June 30, 2026, compared to $0.5 million income for the three months ended June 30, 2025, attributed primarily to an increase in the fair value of the Company’s warrant liability.

Net loss attributable to common stockholders, basic and diluted, for the quarter ended June 30, 2026, was $(1.7) million, or $(0.06) per share of common stock, compared to $(6.5) million loss, or $(0.52) per share of common stock, for the three months ended June 30, 2025. The decreased net loss for the quarter ended June 30, 2026, was attributed primarily to a decrease in the fair value of the Company’s warrant liability.

About Unicycive Therapeutics

Unicycive Therapeutics is a biotechnology company developing novel treatments for kidney diseases. Unicycive’s lead investigational treatment is oxylanthanum carbonate, a novel phosphate binding agent for the treatment of hyperphosphatemia in patients with chronic kidney disease who are on dialysis. Unicycive’s second investigational treatment UNI-494 is intended for the treatment of conditions related to acute kidney injury. It has been granted orphan drug designation (ODD) by the FDA for the prevention of Delayed Graft Function (DGF) in kidney transplant patients and has completed a Phase 1 dose-ranging safety study in healthy volunteers. For more information, please visit Unicycive.com and follow us on LinkedIn and X.

Forward-looking statements

Certain statements in this press release are forward-looking within the meaning of the Private Securities Litigation Reform Act of 1995. These statements may be identified using words such as “anticipate,” “believe,” “forecast,” “estimated” and “intend” or other similar terms or expressions that concern Unicycive’s expectations, strategy, plans or intentions. These forward-looking statements are based on Unicycive’s current expectations and actual results could differ materially. There are several factors that could cause actual events to differ materially from those indicated by such forward-looking statements. These factors include, but are not limited to, clinical trials involve a lengthy and expensive process with an uncertain outcome, and results of earlier studies and trials may not be predictive of future trial results; our clinical trials may be suspended or discontinued due to unexpected side effects or other safety risks that could preclude approval of our product candidates; our dependence on third parties for manufacturing; risks related to business interruptions, which could seriously harm our financial condition and increase our costs and expenses; dependence on key personnel; substantial competition; uncertainties of patent protection and litigation; dependence upon third parties; market acceptance of our products; and risks related to failure to obtain FDA clearances or approvals and noncompliance with FDA regulations. Actual results may differ materially from those indicated by such forward-looking statements as a result of various important factors, including: the uncertainties related to market conditions and other factors described more fully in the section entitled ‘Risk Factors’ in Unicycive’s Annual Report on Form 10-K for the year ended December 31, 2025, and other periodic reports filed with the Securities and Exchange Commission. Any forward-looking statements contained in this press release speak only as of the date hereof, and Unicycive specifically disclaims any obligation to update any forward-looking statement, whether as a result of new information, future events or otherwise.

Investor Contacts:
Kevin Gardner
LifeSci Advisors
[email protected]

Media Contact:
Unicycive Therapeutics
[email protected]

SOURCE: Unicycive Therapeutics, Inc.

View full release here.

Nutriband (NTRB) – Looking Forward To Product Milestones In The Second Half FY2026


Wednesday, August 12, 2026

Robert LeBoyer, Senior Vice President, Equity Research Analyst, Biotechnology, Noble Capital Markets, Inc.

Refer to the full report for the price target, fundamental analysis, and rating.

AVERSA Fentanyl Continues To Make Progress. Nutriband has been working in several areas to advance AVERSA Fentanyl toward the market. These include preparations for the registration trial, manufacturing, and commercialization. We continue to see AVERSA Fentanyl as an important product that could make fentanyl a safe, abuse-resistant option for pain relief.

Clinical Trial Expected Later In FY2026. The AVERSA Fentanyl application for FDA approval requires only a single clinical trial providing data to show that Fentanyl abusers prefer generic patches to the abuse-deterrent AVERSA technology. We expect this to be a short trial with a relatively small number of patients. Manufacturing clinical supplies is progressing, with the trial expected to begin around late Fall 2026.


Get the Full Report

Equity Research is available at no cost to Registered users of Channelchek. Not a Member? Click ‘Join’ to join the Channelchek Community. There is no cost to register, and we never collect credit card information.

This Company Sponsored Research is provided by Noble Capital Markets, Inc., a FINRA and S.E.C. registered broker-dealer (B/D).

*Analyst certification and important disclosures included in the full report. NOTE: investment decisions should not be based upon the content of this research summary. Proper due diligence is required before making any investment decision. 

Release – GeoVax Highlights Gedeptin® Tumor-Priming Strategy as Immuno-Oncology Enters New Phase

GeoVax

Research News and Market Data on GOVX

FDA Approval of Replimune’s Intratumoral Therapy in Combination with Checkpoint Inhibition Reinforces Growing Momentum Behind Tumor-Directed Approaches Designed to Enhance Anti-Tumor Immune Response

ATLANTA, GA – August 11, 2026 – GeoVax Labs, Inc. (Nasdaq: GOVX), a clinical-stage biotechnology company developing vaccines and immunotherapies for infectious diseases and solid tumors, today highlighted the recent U.S. Food and Drug Administration (FDA) accelerated approval of Replimune Group, Inc.’s TUDRIQEV™ (formerly RP1) in combination with nivolumab as an important milestone in the evolution of intratumoral cancer immunotherapy.

The FDA approved TUDRIQEV in combination with nivolumab for adults with unresectable advanced cutaneous melanoma whose disease has progressed following prior anti-PD-1 therapy. The approval followed a favorable FDA Advisory Committee recommendation and represents an important regulatory milestone for localized tumor-directed therapies used in combination with immune checkpoint inhibitors (ICIs).

While TUDRIQEV and GeoVax’s investigational solid tumor therapy Gedeptin® employ different mechanisms of action to achieve tumor cell killing, GeoVax believes the approval reinforces a broader therapeutic paradigm: local destruction of tumor masses may modify the tumor microenvironment and potentially enhance immune responses both within directly treated tumors and at distant tumor sites.

“This approval represents an important milestone that extends well beyond any individual product,” said David A. Dodd, Chairman and Chief Executive Officer of GeoVax. “It provides clinical and regulatory precedent for locally administered therapies used in combination with checkpoint inhibition. We believe this represents an increasingly important direction for cancer immunotherapy.”

Overcoming the “Cold Tumor” Barrier

Earlier this year, Mr. Dodd outlined this emerging concept in an Onco’Zine commentary entitled The Cold Tumor Barrier: Why Promising Oncology Therapies Fail In Vivo – and What It Will Take to Overcome It (Onco’Zine Cold Tumor Barrier). The article describes immunologically “cold” tumors – characterized by limited T-cell infiltration, poor antigen presentation and an immunosuppressive tumor microenvironment – as a fundamental barrier limiting the effectiveness of checkpoint inhibitors and other immunotherapies.

The article argues that durable advances in immuno-oncology may increasingly depend upon therapies capable of priming the tumor microenvironment and converting immunologically “cold” tumors into immune-responsive “hot” tumors, thereby creating the biological conditions necessary for checkpoint inhibitors to function more effectively.

GeoVax believes Gedeptin® represents a differentiated approach to this emerging therapeutic paradigm. Unlike replication-dependent oncolytic viruses, Gedeptin utilizes a gene-directed enzyme prodrug therapy (GDEPT) approach in which a replication-deficient adenoviral vector delivers a bacterial enzyme, purine nucleoside phosphorylase (PNP), directly into tumor tissue. Following administration of fludarabine phosphate, the enzyme converts the inactive prodrug into a potent cytotoxic metabolite within the tumor microenvironment, producing localized tumor destruction and a demonstrated preclinical bystander effect extending beyond directly transduced tumor cells, while promoting anti-tumor immune responses. Importantly, GeoVax believes Gedeptin’s potential therapeutic value extends beyond localized cytotoxicity. 

Published Evidence Supporting Tumor Priming and Checkpoint Inhibitor Synergy

In recently published research in JCI Insight, Gedeptin combined with anti-PD-1 therapy in an immunocompetent preclinical model demonstrated enhanced anti-tumor immune responses, increased CD8+ T-cell infiltration, systemic anti-tumor activity and improved therapeutic outcomes compared with checkpoint inhibition alone.

These findings provide scientific support for the thesis that Gedeptin may function as a tumor-priming immunotherapy, using localized tumor destruction and immune activation to help transform immunologically resistant tumors into tumors more responsive to checkpoint inhibition.

“The objective is not simply to destroy tumor cells locally,” said Kelly McKee, M.D., Chief Medical Officer of GeoVax. “The greater opportunity is to initiate the immune recognition necessary for checkpoint inhibitors to work more effectively. These findings demonstrate Gedeptin’s potential to combine extensive localized tumor killing with immune activation, creating a more favorable tumor microenvironment for checkpoint inhibition.” 

A Converging Immuno-Oncology Strategy

GeoVax believes several recent developments now point toward a converging therapeutic strategy within immuno-oncology:

  • Regulatory precedent: FDA accelerated approval of TUDRIQEV plus nivolumab establishes an approved intratumoral immunotherapy/checkpoint inhibitor combination for patients with advanced melanoma following anti-PD-1 therapy.
  • Scientific evidence: The JCI Insight publication provides preclinical evidence that Gedeptin can enhance anti-tumor immune activity and checkpoint inhibitor responsiveness.
  • Tumor-priming strategy: Growing scientific understanding of the “cold tumor” barrier supports approaches designed to modify the tumor microenvironment before or in conjunction with checkpoint inhibition.

Gedeptin is mechanistically distinct from TUDRIQEV. Rather than relying on viral replication and oncolysis, Gedeptin is designed to generate potent cytotoxic activity directly within the tumor through enzyme-directed prodrug activation, with a substantial bystander effect capable of extending tumor killing beyond cells directly reached by the vector.

GeoVax believes this differentiated mechanism has the potential to position Gedeptin as a tumor-priming immunotherapy designed to enhance the effectiveness of established immunotherapies, rather than simply as another intratumoral therapy. The Company is continuing preparations for the next phase of Gedeptin’s clinical development in combination with pembrolizumab for patients with head and neck squamous cell carcinoma. The planned study is expected to evaluate clinical activity together with biomarkers of immune activation, tumor microenvironment modulation and pathological tumor response.

“The FDA approval of an intratumoral therapy combined with checkpoint inhibition reinforces a therapeutic direction that we believe has significant potential,” concluded Mr. Dodd. “Our objective with Gedeptin is to address one of the fundamental barriers limiting immunotherapy – the inability of immunologically cold tumors to generate an effective anti-tumor immune response. Our published science provides an important foundation for that strategy, and we believe Gedeptin’s differentiated mechanism warrants continued clinical development.” 

About Gedeptin®

Gedeptin® is GeoVax’s investigational gene-directed enzyme prodrug therapy (GDEPT) for the treatment of solid tumors. The therapy utilizes a replication-deficient adenoviral vector to deliver the bacterial enzyme purine nucleoside phosphorylase (PNP) directly into tumors. Following administration of fludarabine phosphate, the PNP enzyme converts the inactive prodrug into a potent cytotoxic metabolite within the tumor microenvironment, producing localized tumor cell death while promoting anti-tumor immune responses. Gedeptin is being developed in combination with immune checkpoint inhibitors for the treatment of solid tumors.

About GeoVax

GeoVax Labs, Inc. is a clinical-stage biotechnology company focused on the development of vaccines and immunotherapies addressing high-consequence infectious diseases and solid tumor cancers. GeoVax’s priority program is GEO-MVA, a Modified Vaccinia Ankara (MVA)–based vaccine targeting mpox and smallpox. The program is advancing under an expedited regulatory pathway, with plans to initiate a pivotal Phase 3 clinical trial in the second half of 2026, to address critical global needs for expanded orthopoxvirus vaccine supply and biodefense preparedness. In oncology, GeoVax is developing Gedeptin®, a gene-directed enzyme prodrug therapy (GDEPT) designed to enhance immune checkpoint inhibitor activity. Gedeptin has completed a multicenter Phase 1/2 clinical trial in advanced head and neck cancer and is being advanced into combination strategies, including planned neoadjuvant and first-line settings. GeoVax maintains a global intellectual property portfolio supporting its infectious disease and oncology programs and continues to evaluate strategic partnerships and funding opportunities aligned with its development priorities. For more information, visit www.geovax.com.

Forward-Looking Statements

This release contains forward-looking statements regarding GeoVax’s business plans. The words “believe,” “look forward to,” “may,” “estimate,” “continue,” “anticipate,” “intend,” “should,” “plan,” “could,” “target,” “potential,” “is likely,” “will,” “expect” and similar expressions, as they relate to us, are intended to identify forward-looking statements. We have based these forward-looking statements largely on our current expectations and projections about future events and financial trends that we believe may affect our financial condition, results of operations, business strategy and financial needs. Actual results may differ materially from those included in these statements due to a variety of factors, including whether: GeoVax is able to obtain acceptable results from ongoing or future clinical trials of its investigational products, GeoVax’s immuno-oncology products and preventative vaccines can provoke the desired responses, and those products or vaccines can be used effectively, GeoVax’s viral vector technology adequately amplifies immune responses to cancer antigens, GeoVax can develop and manufacture its immuno-oncology products and preventative vaccines with the desired characteristics in a timely manner, GeoVax’s immuno-oncology products and preventative vaccines will be safe for human use, GeoVax’s vaccines will effectively prevent targeted infections in humans, GeoVax’s immuno-oncology products and preventative vaccines will receive regulatory approvals necessary to be licensed and marketed, GeoVax raises required capital to complete development, there is development of competitive products that may be more effective or easier to use than GeoVax’s products, GeoVax will be able to enter into favorable manufacturing and distribution agreements, and other factors, over which GeoVax has no control.

Further information on our risk factors is contained in our periodic reports on Form 10-Q and Form 10-K that we have filed and will file with the SEC. Any forward-looking statement made by us herein speaks only as of the date on which it is made. Factors or events that could cause our actual results to differ may emerge from time to time, and it is not possible for us to predict all of them. We undertake no obligation to publicly update any forward-looking statement, whether as a result of new information, future developments or otherwise, except as may be required by law.

Company Contact:

[email protected]

678-384-7220

Media Contact:

Jessica Starman

[email protected] 

Release – Cardiff Oncology Reports Second Quarter 2026 Results and Provides Business Update

Research News and Market Data on CRDF

August 11, 2026

PDF Version

Positive Phase 2 CRDF-004 data presented in an oral session at ASCO support advancement of 30 mg onvansertib plus FOLFIRI/bevacizumab into planned registrational program for first-line RAS-mutated mCRC

Following successful End-of-Phase 2 meeting with FDA, Company aligned on key elements of registrational trial; plans to initiate study in Q1 2027, subject to securing additional financing

Completed $10 million Registered Direct offering, extending cash runway

SAN DIEGO, Aug. 11, 2026 (GLOBE NEWSWIRE) — Cardiff Oncology, Inc. (Nasdaq: CRDF), a clinical-stage biotechnology company leveraging PLK1 inhibition to develop novel cancer therapies, today announced financial results for the second quarter ended June 30, 2026, and provided a business update.

“The second quarter was an important period of progress for Cardiff, highlighted by the presentation of positive Phase 2 data at ASCO and our continued progress in preparation for a planned registrational trial of onvansertib in first-line RAS-mutated metastatic colorectal cancer,” said Mani Mohindru, PhD, President and Chief Executive Officer of Cardiff Oncology. “The updated CRDF-004 results reinforced our confidence in the selected registrational dose and regimen of 30 mg onvansertib in combination with FOLFIRI/bevacizumab. This regimen has demonstrated deep and durable tumor shrinkage over time, reflecting the synergistic mechanisms of action, while maintaining a well-tolerated safety profile with no overlapping or new toxicities when added to standard-of-care therapy.”

Dr. Mohindru continued, “Following our successful End-of-Phase 2 meeting with the FDA, we are preparing to initiate the planned Phase 3 trial in the first quarter of 2027, subject to securing additional financing. We believe the totality of data generated to date strengthens onvansertib’s potential to become an important new treatment option for patients with first-line RAS-mutated metastatic colorectal cancer, an area where there remains significant unmet need.”

Clinical and Regulatory Highlights

Presented Positive Results from Randomized, Controlled Phase 2 CRDF-004 Trial at the 2026 American Society of Clinical Oncology (“ASCO”) Annual Meeting

In June, Cardiff presented positive results from CRDF-004, its ongoing, randomized, controlled, dose-finding Phase 2 clinical trial evaluating onvansertib in combination with standard-of-care (“SoC”) regimens in patients with first-line RAS-mutated metastatic colorectal cancer (“mCRC”), in a rapid oral presentation at the 2026 ASCO Annual Meeting.

The trial achieved its primary goal of selecting the efficacious and safe dose of onvansertib plus SoC regimen for the registrational program. The selected regimen, 30 mg onvansertib in combination with FOLFIRI/bevacizumab (“bev”), demonstrated deep and durable tumor shrinkage, including clinically meaningful improvements in confirmed objective response rate (“ORR”) and progression-free survival (“PFS”) compared to SoC alone, with no additive adverse events observed. Data highlights from the ongoing Phase 2 trial, based on a March 18, 2026 data cut, are listed below, with the full press release available here:

  • The 30 mg onvansertib plus FOLFIRI/bev arm achieved a confirmed ORR of 72.2% compared to 42.1% for FOLFIRI/bev alone, a 30% ORR improvement over SoC. The responses were deeper and more durable in the onvansertib arm.
  • Secondary endpoint of PFS hazard ratio (“HR”) of 0.55 (95% CI: 0.15–2.09) and 0.57 (95% CI: 0.20–1.65) for patients treated with 30 mg onvansertib plus FOLFIRI/bev vs. FOLFIRI/bev by Blinded Independent Central Review (“BICR”) and investigator assessment (“IA”), respectively.
  • Four patients remained on onvansertib treatment beyond 15 months, including two patients beyond 20 months.
  • Onvansertib in combination with SoC regimens continued to be well-tolerated, with no major or unexpected toxicities and no additive adverse events observed.

The Phase 2 trial is still ongoing and as of a June 23, 2026 data cut, 12 patients remain on trial, with 8 patients in the onvansertib (20 or 30 mg) plus FOLFIRI/bev arms and one patient remaining on SoC.

Completed Successful End-of-Phase 2 (“EoP2”) Meeting with FDA and Advanced Phase 3 Readiness Activities

  • Following completion of a successful EoP2 meeting, Cardiff aligned with the FDA on key design elements for its planned registrational Phase 3 trial of onvansertib in first-line RAS-mutated mCRC.
  • The planned randomized, controlled Phase 3 trial is expected to evaluate 30 mg onvansertib in combination with FOLFIRI/bev compared to SoC FOLFIRI/bev as first-line therapy in patients with RAS-mutated mCRC. Cardiff is preparing to initiate the trial in the first quarter of 2027, subject to securing additional financing.

Preclinical Highlights

Presented New Preclinical Data at the 2026 American Association for Cancer Research (“AACR”) Annual Meeting Supporting the Rationale for Onvansertib in Combination with Antibody-Drug Conjugates (“ADCs”)

  • In April, Cardiff presented new preclinical data at the 2026 AACR Annual Meeting supporting the rationale for onvansertib in combination with ADCs. The data demonstrated that onvansertib enhanced the activity of the HER2-targeted antibody-drug conjugate trastuzumab deruxtecan, driving tumor regression and overcoming resistance in HER2-low breast cancer models.

Corporate Update

  • In February 2026, the Company received written notice from its licensor, Nerviano Medical Sciences S.r.l. (“NMS”), alleging that the Company was in material breach of the license agreement. NMS subsequently purported to terminate the license agreement based on the Company’s alleged material breach. The Company filed a lawsuit in May 2026 in the U.S. District Court for the Southern District of California seeking a declaratory judgment that it is not in material breach and injunctive relief requiring NMS to continue performing under the license agreement. The Company believes that NMS’s purported termination is legally ineffective, factually unsupported and procedurally improper, and the Company plans to continue performing under the license agreement.
  • In July, Cardiff announced a $10 million registered direct offering of common stock and warrants to support working capital and general corporate purposes. The full press release is available here.

Second Quarter 2026 Financial Results

Liquidity, cash burn, and cash runway

As of June 30, 2026, Cardiff Oncology had approximately $34.5 million in cash, cash equivalents, and short-term investments. The amount as of June 30, 2026 does not include proceeds from the registered direct offering completed subsequent to quarter end.

Net cash used in operating activities for the six months ended June 30, 2026 was approximately $24.1 million, an increase of $3.0 million from $21.1 million for the same period in 2025.

Based on its current expectations and projections, the Company believes its current cash resources are sufficient to fund its operations into the third quarter of 2027.

Operating results

Total operating expenses were approximately $22.6 million for the six months ended June 30, 2026, a decrease of $6.8 million from $29.4 million for the same period in 2025. The decrease in operating expenses was primarily due to a decrease of $9.4 million in R&D expenses, mainly related to the completion of clinical trials, as well as fewer patients still on treatment in the Phase 2 mCRC trial, and a reduction in preclinical activities as the Company focuses on its upcoming Phase 3 mCRC trial. The decrease in expenses was partially offset by an increase of $2.6 million in SG&A expenses, primarily for employee severance agreements and corresponding modifications of stock options, as well as an increase in attorney costs related to Cardiff Oncology’s ongoing licensing dispute.

About Cardiff Oncology, Inc.

Cardiff Oncology is a clinical-stage biotechnology company advancing innovative cancer treatments focused on PLK1 inhibition, a validated oncology target with practice-changing potential. Our lead asset, onvansertib, is a highly specific, oral PLK1 inhibitor currently being evaluated in a Phase 2 trial for first-line treatment of RAS-mutated metastatic colorectal cancer (“mCRC”), addressing a large, underserved patient population with high unmet need. Onvansertib is also under investigation in other PLK1-driven cancers through ongoing investigator-initiated trials and has shown robust single-agent clinical activity in hard-to-treat tumors. By targeting tumor vulnerabilities, we aim to overcome treatment resistance and deliver improved clinical outcomes for patients.

For more information, please visit https://www.cardiffoncology.com.

Forward-Looking Statements

Certain statements in this press release are forward-looking within the meaning of the Private Securities Litigation Reform Act of 1995. These statements may be identified using words such as “anticipate,” “believe,” “forecast,” “estimated” and “intend” or other similar terms or expressions that concern Cardiff Oncology’s expectations, strategy, plans or intentions. These forward-looking statements are based on Cardiff Oncology’s current expectations and actual results could differ materially. There are several factors that could cause actual events to differ materially from those indicated by such forward-looking statements. These factors include, but are not limited to, clinical trials involve a lengthy and expensive process with an uncertain outcome, and results of earlier studies and trials may not be predictive of future trial results; our clinical trials may be suspended or discontinued due to unexpected side effects or other safety risks that could preclude approval of our product candidate; results of preclinical studies or clinical trials for our product candidate could be unfavorable or delayed; our need for additional financing; uncertainty as to the outcome of pending litigation against Nerviano Medical Sciences S.r.l. (NMS) with respect to our license agreement with NMS; risks related to business interruptions, including the outbreak of COVID-19 coronavirus and cyber-attacks on our information technology infrastructure, which could seriously harm our financial condition and increase our costs and expenses; uncertainties of government or third-party payer reimbursement; dependence on key personnel; limited experience in marketing and sales; substantial competition; uncertainties of patent protection and litigation; dependence upon third parties; and risks related to failure to obtain FDA clearances or approvals and noncompliance with FDA regulations. There are no guarantees that our product candidate will be utilized or prove to be commercially successful. Additionally, there are no guarantees that future clinical trials will be completed or successful or that our product candidate will receive regulatory approval for any indication or prove to be commercially successful. Investors should read the risk factors set forth in Cardiff Oncology’s Form 10-K for the year ended December 31, 2025, and other periodic reports filed with the Securities and Exchange Commission. While the list of factors presented here is considered representative, no such list should be considered to be a complete statement of all potential risks and uncertainties. Unlisted factors may present significant additional obstacles to the realization of forward-looking statements. Forward-looking statements included herein are made as of the date hereof, and Cardiff Oncology does not undertake any obligation to update publicly such statements to reflect subsequent events or circumstances.

Investor Contact:
Candice Masse
astr partners
[email protected]

Media Contact:
Amy Bonanno
Lyra Strategic Advisory
[email protected]

View full release here.

NeuroSense Therapeutics Ltd. (NRSN) – NeuroSense Announces Target Date For Canadian PrimeC Approval Application


Tuesday, August 11, 2026

Robert LeBoyer, Senior Vice President, Equity Research Analyst, Biotechnology, Noble Capital Markets, Inc.

Refer to the full report for the price target, fundamental analysis, and rating.

Pre-Application Process Has Been Completed. NeuroSense announced that it has completed Pre-NDS meetings with Health Canada and plans to submit a New Drug Submission (NDS) for PrimeC in its ALS indication. These meetings focused on whether the data could support approval and the submission requirements. The target date is December 2026. We see this as good news that is consistent with our expectations.

We View The Canadian NDS Process As An Important Milestone For PrimeC. The NDS application will include the Phase 2b PARADIGM trial data, with additional preclinical and supporting data. The primary endpoint in the trial showed a reduction in TDP-43 (TAR DNA-binding Protein 43, a protein that drives ALS progression and deterioration). The data also showed increased median survival, improved functional assessments, biomarkers showing slower disease progression, as well as safety and tolerability. Approval would be based on Health Canada’s analysis of these data.


Get the Full Report

Equity Research is available at no cost to Registered users of Channelchek. Not a Member? Click ‘Join’ to join the Channelchek Community. There is no cost to register, and we never collect credit card information.

This Company Sponsored Research is provided by Noble Capital Markets, Inc., a FINRA and S.E.C. registered broker-dealer (B/D).

*Analyst certification and important disclosures included in the full report. NOTE: investment decisions should not be based upon the content of this research summary. Proper due diligence is required before making any investment decision. 

NanoViricides (NNVC) – NanoViricides Receives Regulatory Approval To Begin Phase 2 For Ebola In Africa


Tuesday, August 11, 2026

Robert LeBoyer, Senior Vice President, Equity Research Analyst, Biotechnology, Noble Capital Markets, Inc.

Refer to the full report for the price target, fundamental analysis, and rating.

Regulatory Approval Allows the Phase 2 Trial for NV-387 To Begin. NanoViricides has received approval to proceed with its Phase 2 trial of NV-387 for the treatment of Ebola in the Democratic Republic of Congo (DRC). We expect the Ebola trial to be followed by a separate Phase 2 trial in Mpox, also to be conducted in the DRC. This is consistent with our expected time frame for the trials.

Previous Preparations Should Allow Treatment To Start Soon. NanoViricides has completed delivery of clinical supplies of NV-387 oral solid formulation (gummies) for treatment of the trial. The trial will be conducted by OM Sai Clinical Research, a contract research organization (CRO) based in India. The CRO has assembled a clinical team with a Principal Investigator, local clinicians, and a university in the region to support the trial. The approval by ACOREP (Autorité Congolaise de Réglementation Pharmaceutique, the Congolese Pharmaceutical Regulatory Authority) should allow the trial to start patient treatment shortly.


Get the Full Report

Equity Research is available at no cost to Registered users of Channelchek. Not a Member? Click ‘Join’ to join the Channelchek Community. There is no cost to register, and we never collect credit card information.

This Research is provided by Noble Capital Markets, Inc., a FINRA and S.E.C. registered broker-dealer (B/D).

*Analyst certification and important disclosures included in the full report. NOTE: investment decisions should not be based upon the content of this research summary. Proper due diligence is required before making any investment decision. 

Release – NanoViricides Has Received Regulatory Approval for a Phase II Clinical Trial of NV-387 Oral Gummies as a Treatment for Ebola in DR Congo To Fight the Current Largest Ever and Expanding Ebola Outbreak

NanoViricides Has Received Regulatory Approval for a Phase II Clinical Trial of NV-387 Oral Gummies as a Treatment for Ebola in DR Congo To Fight the Current Largest Ever and Expanding Ebola Outbreak

Research News and Market Data on NNVC

Monday, 10 August 2026 08:30 AM

SHELTON, CT / ACCESS Newswire / August 10, 2026 / NanoViricides, Inc. (NYSE American:NNVC) (the “Company”), a clinical stage leader developing antiviral drugs that viruses cannot escape, announces that it has received regulatory approval to begin a Phase II Clinical Trial of NV-387 Oral Gummies as a Treatment for the Current Bundibugyo Ebolavirus and other Ebola viruses, from the local regulatory agency ACOREP in the Democratic Republic of Congo (DRC).

NanoViricides has retained Om Sai Clinical Research Private Limited, India, (Om Sai CRO) as the CRO for this Phase II clinical trial for Ebola in DRC. Om Sai CRO has been instrumental in putting together a team with a renowned Principal Investigator and other renowned experts and with support from a well known University in the Ebola-affected region to lead and execute the clinical trial of NV-387 Oral Gummies as a Treatment for Ebola viruses in DRC.

The Principal Investigator (PI) sent in the application for the clinical trial to ACOREP which has now been approved. The PI, with the DRC team, has already been performing initial preparations in anticipation of the clinical trial approval, in order to speed up the date to the first dosing of patients.

“We believe NanoViricides is well positioned to provide an Oral Ebola treatment to save lives with our NV-387 Oral Gummies drug product that is already in place in DRC,” said Anil R. Diwan, PhD, President of the Company, adding, “This unique and revolutionary oral broad-spectrum antiviral drug will now be tested in a clinical trial, and we thank all involved in making this possible.”

The current Ebola Virus Disease (EVD) caused by the Bundibogyo ebolavirus (BDBV) is now the largest ever ebola outbreak in DRC. As of August 6, 2026, there have been 4,141 confirmed cases and 1,889 confirmed deaths due to this virus. Only 829 confirmed cases have been reported to have recovered from the disease. The crude fatality rate (crude CFR) is 46%, according to the WHO1. The outbreak continues to grow, and during the most recent complete reporting week, the highest weekly number of reported cases (567) and deaths (296) to date were recorded, according to the WHO2. Reportedly, “(it) is spreading like a wildfire”3.

This Ebola outbreak continues to increase in spread and is now present in at least five provinces in DRC4. More concerning is the fact that over 80% of new cases are outside of known contact lists, leading to the projection that the extent of the outbreak is at least two times or more larger than the reported confirmed cases5. Additionally, Ebola is now found to have spread into displacement camps that host over 4.4 million displaced persons due to internal warfare, adding another high risk population pool with poor drinking water, sanitation and medical resources to further fuel this outbreak, according to the UN New Service6.

There is thus a tremendous urgency to validate a drug that works against this ebolavirus in short and decisive clinical trials for minimizing further spread by treating patients and for saving lives. Om Sai CRO, in consultation with renowned scientists in DRC, has designed the Phase II clinical trial with this particular objective.

In contrast, the PARTNERS clinical trial (see below) will require over 1,000 patients to be treated and may not yield results for at least more than a year. A similar large collaborative clinical trial effort in the West Africa outbreak resulted in US FDA approval of two antibody drugs only specifically for EBOV Zaire, which are not deemed to be useful in the current outbreak without further clinical trials.

NV-387 is the only orally active agent under consideration for clinical trial as a treatment of Ebola to the best of our knowledge. In an epidemic scenario in resource limited settings such as in DRC, we believe an oral drug is a highly advantageous feature.

Other treatments under consideration require infusions. Infusions are difficult to implement and also are not scalable in a large outbreak scenario such as this Ebola virus outbreak if it continues to grow, as has been widely expected.

A clinical trial, called the “PARTNERS” clinical trial, evaluating Remdesivir infusion, an antibody cocktail MBP134 infusion, and MBP134 infusion plus Remdesivir infusion, has started according to WHO with first patient having received infusion of the antibody cocktail on July 2, 20267.

“Although this antiviral (Remdesivir) proved to be ineffective at targeting the Zaire Ebolavirus, there remains hope that it could have some benefit against the Bundibugyo virus, particularly if used in combination with MBP-134,” according to an article in Forbes explaining the “PARTNERS” clinical trial by the WHO organized collaboration8. The article also notes that MBP134 contains two separate antibodies designed to, taken together, recognize multiple Ebola species.

Antibodies are highly specific to a particular strain of the virus and usually are not very effective against variants of the same virus that arise in the field. Viruses also escape antibodies readily by mutations in the field.

The Company notes that NV-387 was previously found to be superior to Remdesivir in a lethal animal model of a viral disease. The Company believes this superiority of NV-387 is reasonably expected to extend to the current novel Bundibugyo ebolavirus strain.

There is no approved Treatment or Vaccine for the new variant of the Bundibugyo Ebolavirus (BDBV) that is causing the current rapidly expanding outbreak of the Ebolavirus Disease (EVD) in DRC. The rare Bundibugyo strain of Ebola virus causing the current outbreak appears to be its new variant, likely freshly introduced from some animal source9, such as fruit bats.

A new clinical trial of an Oxford University designed Bundibugyo-specific vaccine has also started in DRC, in addition to the treatment trials of antibody and remdesivir infusions (ibid, #8). Further, Ervebo, a vaccine approved for Ebolavirus Zaire, may also enter clinical trials for the protection of uninfected persons from the distinctly different Bundibugyo ebolavirus due to the enormity of the emergency posed by the current outbreak despite reservations regarding a potentially imperfect vaccine10.

Om Sai is the CRO leading the Company’s Phase II clinical trial of NV-387 Oral Gummies as a Treatment for Mpox in DRC, and the same CRO is also leading the newly approved Ebola clinical trial.

Sufficient quantity of NV-387 Oral Gummies Drug Product for starting the clinical trial against Ebola is already available in DRC. This drug product was shipped to DRC for the ensuing Phase II clinical trial of NV-387 for the Treatment of Mpox and also to support the Phase II clinical trial for the Treatment of Ebola only upon approval by the regulatory agency, which has now been approved.

“We believe NV-387 could be revolutionary in this fight against Ebola, if it is found to be effective,” said Anil R. Diwan, PhD, adding, “It is an oral drug, in contrast to others that are infusions. Thus evaluating if NV-387 treatment works is of paramount importance to combat this and future Ebola and Marburg outbreaks.”

NV-387 is a broad-spectrum antiviral that mimics the host-side features that the virus requires, and is likely to be effective against Ebola viruses because they use the same host-side feature mimicked by NV-387.

It is highly unlikely that viruses can escape NV-387, because this drug mimics the features on host cells that the viruses continue to require even as they mutate or evolve in the field.

Additionally, NV-387 Oral Gummies is a drug product readily delivered orally. It does not even require swallowing effort or water, because it dissolves in the mouth by itself, simplifying delivery for even sick individuals with swallowing difficulties.

This oral delivery is an important feature that puts NV-387, a broad-spectrum antiviral, as being superior to the other approaches.

“Only safe and effective broad-spectrum antiviral drugs like NV-387 that can effectively tackle most viral infections will enable the world to combat viruses and defend the global population in the war against known and unknown nanoscopic enemies that are viruses,” commented Dr. Diwan, adding, “Today, NV-387 is the only drug in clinical development with such broad-spectrum potential that promises to combat diverse epidemics like Mpox and Ebola, to the best of our knowledge.”

While there is currently minimal risk of Ebola in the USA, the CDC’s mathematical models suggested this Central African outbreak could grow to 10,000 to 20,000 cases and 2,000 to 4,000 deaths within just three months, rivaling the largest outbreak to date in 2014-201611. Unfortunately, the outbreak appears to be even more aggressive than the CDC model, with close to 2,000 deaths in less than three months, over 4,000 confirmed cases, and over 10,000 estimated total cases12.

The outbreak which was declared a Public Health Emergency of International Concern (“PHEIC”) by the WHO on May 17, 2026, continues to rapidly expand, outpacing containment efforts. The outbreak arose in a high traffic region bordering the Democratic Republic of Congo (DRC), with travel contacts to Uganda, and South Sudan and with 11 more nations in Africa at risk13.

NV-387 is a broad-spectrum antiviral that mimics the host-side feature called heparan sulfate proteoglycan (HSPG) that over 90-95% of human pathogenic viruses require for infecting cells. No matter how much the virus changes in the field, it continues to use HSPG, and therefore it cannot escape the drug NV-387. In contrast, Remdesivir is a small molecule inhibitor of the viral RDRP enzyme needed for making copies of the viral genome, and the virus can possibly escape by small number of mutations.

All Ebola viruses utilize HSPG as the attachment receptor prior to gaining entry into the cell. Thereafter, followed by entry into the cell inside endosomes, the ebolavirus surface glycoprotein is substantially degraded, opening up its site for binding to its cognate receptor called NPC1, thereby entering into the cytoplasm where the next steps in its replication begin.

Thus there is a strong rationale that NV-387 could be highly effective against Ebola virus infections, not just Bundibugyo, but also the Sudan and other viruses for which there are no treatments.

NV-387 is available as an oral medication that has excellent stability at room temperature, enabling ease of transport, distribution, and delivery to patient. NV-387 oral gummies dissolve naturally in the mouth and do not require tablet swallowing, which is difficult for children, seniors, and also patients with sore throat.

All previous anti-Ebola efforts have been focused on vaccines and antibodies14. This has led to approval of therapies that are specific to the Ebolavirus Zaire strain only, albeit with limited effectiveness. This leaves out all other filoviruses of consequence: Sudan, Marburg, and the more rare Bundibugyo with no treatment or vaccine.

In contrast, if NV-387, as a broad-spectrum antiviral, is found to be effective against the Bundibugyo virus, it will likely be effective against all ebolaviruses and possibly all filoviruses; that would be a game changer for pandemic preparedness.

The case fatality rate of ebolaviruses has generally been approximately 50% in recent outbreaks, with improvements in care, including hydration therapy, corticosteroids, and other usual symptomatic treatments. Ebola viruses spread via bodily fluid secretions including fomites/sputum, as well as semen/genital secretions. Ebola virus can remain in survivors even as many as 965 days after the disease without symptoms, and can transmit through bodily secretions, suggesting possible latency. Many recent outbreaks have been ignited as a result of such reawakened-transmitted virus from a survivor. Sexual transmission was documented even as late as 482 days after disease. This persistence and possible latency of ebolavirus in immune-privileged organs (e.g. brain, eyes, gonads, where antibodies are not operative) makes it a uniquely serious threat for global transmission and sustained outbreaks.

So far, BDBV has demonstrated variable CFR ranging from under 15% (in Uganda, 2026), to 46% (in DRC, based on current confirmed cases and fatalities numbers, as of August 6, 2026). Therefore, BDBV is of great concern as a potential pandemic disease. However, it is believed that ebolaviruses do not transmit via respiratory droplets or aerosols and rather require extensive contact with bodily fluids of an infected person. In addition, within DRC and internationally, certain protective quarantine measures for travel from the outbreak areas have been implemented. Therefore, currently there is no apparent threat of a global pandemic.

An irony is that because of the high case fatality rate (CFR) approaching 50%, the spread of ebolaviruses remains rather limited. If a variant emerges with a reduced CFR, say in the range of 5-15%, the potential threat of global pandemic from such an outbreak would increase substantially.

With ever-increasing global travel, local outbreaks such as ebola can quickly travel far and wide potentially causing global pandemics, as was the case with COVID-19, if not caught in time. It is not feasible to produce a new vaccine and a new set of antibody drugs to combat every possible virus. Even if vaccines and antibodies are produced, the virus would escape by generating variants, as the world has witnessed during the COVID-19 pandemic.

The US Government is active in ensuring that suspected or confirmed ebolavirus cases do not enter the general population in the USA. To this end, travel from DRC has been restricted, with pre-travel quarantine requirements imposed, and suspect travelers are directed to screening at specific airports and may be further quarantined.

ABOUT NANOVIRICIDES

NanoViricides, Inc. (the “Company”) (www.nanoviricides.com) is a clinical stage company that is creating special purpose nanomaterials for antiviral therapy. The Company’s novel nanoviricide™ class of drug candidates and the nanoviricide™ technology are based on intellectual property, technology and proprietary know-how of TheraCour Pharma, Inc. The Company has a Memorandum of Understanding with TheraCour for the development of drugs based on these technologies for all antiviral infections. The MoU does not include cancer and similar diseases that may have viral origin but require different kinds of treatments.

The Company has obtained broad, exclusive, sub-licensable, field licenses to drugs developed in several licensed fields from TheraCour Pharma, Inc. The Company’s business model is based on licensing technology from TheraCour Pharma Inc. for specific application verticals of specific viruses, as established at its foundation in 2005.

Our lead drug candidate is NV-387, a broad-spectrum antiviral drug that we plan to develop as a treatment of RSV, COVID, Long COVID, Influenza, and other respiratory viral infections, as well as MPOX/Smallpox infections. Our other advanced drug candidate is NV-HHV-1 for the treatment of Shingles. The Company cannot project an exact date for filing an IND for any of its drugs because of dependence on a number of external collaborators and consultants. The Company is currently focused on advancing NV-387 into Phase II human clinical trials.

NV-CoV-2 (API NV-387) is our nanoviricide drug candidate for COVID-19 that does not encapsulate remdesivir. NV-CoV-2-R is our other drug candidate for COVID-19 that is made up of NV-387 with remdesivir encapsulated within its polymeric micelles. The Company believes that since remdesivir is already US FDA approved, our drug candidate encapsulating remdesivir is likely to be an approvable drug, if safety is comparable. Remdesivir is developed by Gilead. The Company has developed both of its own drug candidates NV-CoV-2 and NV-CoV-2-R independently.

The Company is also developing drugs against a number of viral diseases including oral and genital Herpes, viral diseases of the eye including EKC and herpes keratitis, H1N1 swine flu, H5N1 bird flu, seasonal Influenza, HIV, Hepatitis C, Rabies, Dengue fever, and Ebola virus, among others. NanoViricides’ platform technology and programs are based on the TheraCour® nanomedicine technology of TheraCour, which TheraCour licenses from AllExcel. NanoViricides holds a worldwide exclusive perpetual license to this technology for several drugs with specific targeting mechanisms in perpetuity for the treatment of the following human viral diseases: Human Immunodeficiency Virus (HIV/AIDS), Hepatitis B Virus (HBV), Hepatitis C Virus (HCV), Rabies, Herpes Simplex Virus (HSV-1 and HSV-2), Varicella-Zoster Virus (VZV), Influenza and Asian Bird Flu Virus, Dengue viruses, Japanese Encephalitis virus, West Nile Virus, Ebola/Marburg viruses, and certain Coronaviruses. The Company intends to obtain a license for RSV, Poxviruses, and/or Enteroviruses if the initial research is successful. As is customary, the Company must state the risk factor that the path to typical drug development of any pharmaceutical product is extremely lengthy and requires substantial capital. As with any drug development efforts by any company, there can be no assurance at this time that any of the Company’s pharmaceutical candidates would show sufficient effectiveness and safety for human clinical development. Further, there can be no assurance at this time that successful results against coronavirus in our lab will lead to successful clinical trials or a successful pharmaceutical product.

This press release contains forward-looking statements that reflect the Company’s current expectation regarding future events. Actual events could differ materially and substantially from those projected herein and depend on a number of factors. Certain statements in this release, and other written or oral statements made by NanoViricides, Inc. are “forward-looking statements” within the meaning of Section 27A of the Securities Act of 1933 and Section 21E of the Securities Exchange Act of 1934. You should not place undue reliance on forward-looking statements since they involve known and unknown risks, uncertainties and other factors which are, in some cases, beyond the Company’s control and which could, and likely will, materially affect actual results, levels of activity, performance or achievements. The Company assumes no obligation to publicly update or revise these forward-looking statements for any reason, or to update the reasons actual results could differ materially from those anticipated in these forward-looking statements, even if new information becomes available in the future. Important factors that could cause actual results to differ materially from the company’s expectations include, but are not limited to, those factors that are disclosed under the heading “Risk Factors” and elsewhere in documents filed by the company from time to time with the United States Securities and Exchange Commission and other regulatory authorities. Although it is not possible to predict or identify all such factors, they may include the following: demonstration and proof of principle in preclinical trials that a nanoviricide is safe and effective; successful development of our product candidates; our ability to seek and obtain regulatory approvals, including with respect to the indications we are seeking; the successful commercialization of our product candidates; and market acceptance of our products.

The phrases “safety”, “effectiveness” and equivalent phrases as used in this press release refer to research findings including clinical trials as the customary research usage and do not indicate evaluation of safety or effectiveness by the US FDA.

FDA refers to US Food and Drug Administration. IND application refers to “Investigational New Drug” application. cGMP refers to current Good Manufacturing Practices. CMC refers to “Chemistry, Manufacture, and Controls”. CHMP refers to the Committee for Medicinal Products for Human Use, which is the European Medicines Agency’s (EMA) committee responsible for human medicines. API stands for “Active Pharmaceutical Ingredient”. WHO is the World Health Organization. R&D refers to Research and Development.

Contact:
NanoViricides, Inc.
[email protected]

Public Relations Contact:
[email protected]

1 https://www.who.int/emergencies/alert-and-response. The crude CFR is estimated as number of deaths (1,889) divided by number of reported confirmed cases (4,141) since May 16, the declaration of the epidemic. Most of the remaining 1,423 cases are likely to be continuing as disease-stricken. The crude CFR does not take into account the delay period between case confirmation to case fatality (i.e. the disease duration), and results in an underestimate of the case fatality rate while the epidemic is still increasing, as is the case at present in DRC.

The largest ever Ebola outbreak was the West Africa Ebolavirus (Zaire) outbreak, in 2013-2016, that killed more than 11,000 people out of at least 28,000 cases. The current outbreak, caused by a different, rare Bundibugyo strain of the ebolavirus, has already surpassed that outbreak in becoming the fastest growing Ebola outbreak to date, according to the WHO.

2 https://www.who.int/emergencies/disease-outbreak-news/item/2026-DON614.

3 https://www.aljazeera.com/news/2026/7/25/ebola-deaths-in-drc-surge-past-1300-as-virus-spreading-like-a-wildfire

4 https://www.aljazeera.com/news/2026/7/20/ebola-death-toll-in-drc-surges-to-at-least-930-as-outbreak-gathers-pace

https://www.aljazeera.com/news/2026/7/16/ebola-spreading-more-quickly-in-drc-while-uganda-is-close-to-being-virus-free

5 https://www.msn.com/en-us/health/other/congos-ebola-outbreak-spreads-to-two-more-provinces/ar-AA27NkjT

6 https://www.msn.com/en-xl/africa/top-stories/ebola-virus-reaches-displacement-camps-in-dr-congo/ar-AA29A5ts?ocid=BingNewsSerp

7 https://www.reuters.com/business/healthcare-pharmaceuticals/trial-bundibugyo-ebola-treatment-starts-drc-who-says-2026-07-02/

8 https://www.forbes.com/sites/omerawan/2026/07/07/new-clinical-trials-offer-hope-in-the-fight-against-ebola-in-the-democratic-republic-of-congo/

9 https://virological.org/t/initial-genomes-from-may-2026-bundibugyo-virus-disease-outbreak-in-the-democratic-republic-of-the-congo-and-uganda/1032

10 https://www.msn.com/en-us/health/general/ebola-cases-top-4-000-in-drc-as-who-urges-ervebo-vaccine-trial/ar-AA29CX9f?ocid=BingNewsSerp .

11 https://www.cdc.gov/media/releases/2026/update-on-ebola-outbreak-in-the-democratic-republic-of-the-congo-and-uganda-6-5-2026.html

12 The WHO and Africa CDC have estimated that the confirmed case number substantially under-represents actual case numbers which could be at least double or even more than confirmed cases. See #5.

13 https://www.forbes.com/sites/maryroeloffs/2026/05/25/african-health-officials-on-ebola-this-is-too-much-live-updates/

14 Substantial work was also performed to develop small chemical potentially broad-spectrum agents. Remdesivir was the only small chemical that entered the PALM clinical trials ca. 2018-2019 but failed to show effectiveness. Small chemicals are readily escaped by viruses often with just single mutations.

SOURCE: NanoViricides
Release – Greenwich LifeSciences Extends Lock-up of Directors and Officers to January 31, 2027

Greenwich LifeSciences

Research News and Market Data on GLSI

 Download as PDFAugust 10, 2026 6:00am EDT

STAFFORD, Texas, Aug. 10, 2026 (GLOBE NEWSWIRE) — Greenwich LifeSciences, Inc. (Nasdaq: GLSI) (the “Company”), a clinical-stage biopharmaceutical company focused on its Phase III clinical trial, FLAMINGO-01, which is evaluating Fast Track designated GLSI-100, an immunotherapy to prevent breast cancer recurrences, today announced that its Board of Directors has extended the lock-up of the shares owned by the Company’s directors, officers, and existing pre-IPO investors to January 31, 2027 which is approximately 76 months from the date of the Company’s IPO. During this period, current officers, directors and certain shareholders will not be able to sell their shares of the Company’s common stock unless otherwise modified by the Board of Directors. After January 31, 2027, the quantity of these locked-up shares that can be sold daily and over various periods of time will be restricted under a leak-out plan unless otherwise modified by the Board of Directors.

The Board could choose to end all or a small percentage of the 100% lock-up at any time for any reason, including prior to any data announcements, interim analyses, strategic transactions, such as an acquisition or partnership, or financial transactions, such as royalty or strategic investor transactions that provide a potential bridge to commercialization. The Board could also choose to implement a pre-determined leak-out plan and/or a 10b5-1 selling program at any time for any reason that allows for the organized independent selling of some of the locked-up shares by a third party over a specified period of time. Such decisions may or may not be announced at the time of the board’s decision.

CEO Snehal Patel commented, “We attended and presented at the BIO conference in June 2026 and plan to attend the BIO-Europe conference later this year, where strategic and financing transactions can be originated or concluded. With the further derisking of the Phase III clinical trial, more than doubling of enrollment and event rate, interest from leading clinicians and hospitals in up to 200 sites in the US and Europe, screening of over 1,500 patients, and the transformation of the interim analysis into a big pharma like study similar in design to recent large breast cancer studies, we are expecting a continued up-tick in the quantity and seriousness of such discussions.”

About Greenwich LifeSciences, Inc.

Greenwich LifeSciences is a clinical-stage biopharmaceutical company focused on the development of GP2, an immunotherapy to prevent breast cancer recurrences in patients who have previously undergone surgery. GP2 is a 9 amino acid transmembrane peptide of the HER2 protein, a cell surface receptor protein that is expressed in a variety of common cancers, including expression in 75% of breast cancers at low (1+), intermediate (2+), and high (3+ or over-expressor) levels. Greenwich LifeSciences has commenced a Phase III clinical trial, FLAMINGO-01. For more information on Greenwich LifeSciences, please visit the Company’s website at www.greenwichlifesciences.com and follow the Company’s Twitter at https://twitter.com/GreenwichLS.

Forward-Looking Statement Disclaimer

Statements in this press release contain “forward-looking statements” that are subject to substantial risks and uncertainties. All statements, other than statements of historical fact, contained in this press release are forward-looking statements. Forward-looking statements contained in this press release may be identified by the use of words such as “anticipate,” “believe,” “contemplate,” “could,” “estimate,” “expect,” “intend,” “seek,” “may,” “might,” “plan,” “potential,” “predict,” “project,” “target,” “aim,” “should,” “will,” “would,” or the negative of these words or other similar expressions, although not all forward-looking statements contain these words. Forward-looking statements are based on Greenwich LifeSciences Inc.’s current expectations and are subject to inherent uncertainties, risks and assumptions that are difficult to predict, including statements regarding the intended use of net proceeds from the public offering; consequently, actual results may differ materially from those expressed or implied by such forward-looking statements. Further, certain forward-looking statements are based on assumptions as to future events that may not prove to be accurate. These and other risks and uncertainties are described more fully in the section entitled “Risk Factors” in Greenwich LifeSciences’ Annual Report on the most recent Form 10-K for the year ended December 31, 2025, and other periodic reports filed with the Securities and Exchange Commission. Forward-looking statements contained in this announcement are made as of this date, and Greenwich LifeSciences, Inc. undertakes no duty to update such information except as required under applicable law.

Company Contact
Snehal Patel
Investor Relations
Office: (832) 819-3232
Email: [email protected]

Investor & Public Relations Contact for Greenwich LifeSciences
Dave Gentry
RedChip Companies Inc.
Office: 1-800-RED CHIP (733 2447)
Email: [email protected]

Primary Logo

Source: Greenwich LifeSciences, Inc.

Released August 10, 2026